FDA Advisory Committee Recommends Adding Six Peptides to Compounding Bulks List in Landmark Regulatory Shift

The landscape of modern regenerative and functional medicine faces a potential turning point following a pivotal two-day meeting by the United States Food and Drug Administration’s (FDA) Pharmacy Compounding Advisory Committee (PCAC). Convened on July 23 and 24, the advisory panel evaluated whether a specific cohort of peptide substances should be integrated into the 503A Bulks List under the Federal Food, Drug, and Cosmetic Act. A favorable placement on this list would grant licensed compounding pharmacies the legal framework to prepare these formulations based on individual physician prescriptions, effectively steering patients away from unregulated online procurement channels.
In a departure from historically restrictive precedents, the committee voted to recommend the inclusion of six prominent peptides: BPC-157, MOTS-c, KPV, TB-500, Epitalon, and Semax. While these advisory votes do not constitute immediate legislative or regulatory changes, they signal a marked ideological shift within federal oversight regarding how emerging therapeutic peptides are evaluated. The development brings to the forefront a broader national conversation concerning health freedom, clinical standardization, and the urgent need for structured medical infrastructure to support rapidly expanding therapeutic markets.
Background and Regulatory Context
To understand the weight of the recent PCAC decision, it is necessary to examine the regulatory paradox surrounding peptide therapies in the United States. Over the past decade, consumer and clinical interest in bioactive peptides—short chains of amino acids that act as signaling molecules in the body—has escalated dramatically. Proponents of peptide medicine highlight their potential in tissue repair, mitochondrial optimization, immune modulation, and cognitive enhancement.

However, because many of these substances exist in a regulatory gray zone, they have largely remained inaccessible through traditional pharmaceutical channels. Under Section 503A of the Federal Food, Drug, and Cosmetic Act, traditional compounding pharmacies are generally restricted from compounding substances that are essentially copies of commercially available drugs or those that present significant safety concerns. When the FDA fails to clarify the status of bulk drug substances, clinicians and patients find themselves navigating a precarious landscape. Advocates frequently warn that strict regulatory prohibitions do not eliminate consumer demand; rather, they drive vulnerable patients toward unverified overseas vendors and illicit online sources, raising legitimate safety, purity, and dosing concerns.
The July 2026 PCAC hearings were designed to assess whether scientific data justified bringing certain heavily utilized peptides into regulated compounding channels. The advisory body’s affirmative recommendations for BPC-157, MOTS-c, KPV, TB-500, Epitalon, and Semax represent the first major institutional crack in a previously rigid defensive posture maintained by federal regulators.
Chronology and Next Steps in the Rulemaking Process
Despite widespread celebration across holistic and functional medicine communities, legal experts and public health analysts emphasize that the PCAC vote is strictly advisory. The formal rule-making process requires substantial time, bureaucratic review, and administrative clearance before any policy changes are codified into federal law.
The anticipated timeline for formal adoption typically spans well over a year. Following the committee’s vote, the FDA must review the panel’s findings, conduct internal safety and risk assessments, and publish a proposed rule in the Federal Register. This phase is traditionally followed by a public comment period, during which manufacturers, clinical associations, and patients can submit data or objections. Only after synthesizing this feedback will the agency issue a final determination. Consequently, none of the six peptides are immediately legal to compound, and prescribers must continue to exercise clinical judgment while monitoring federal updates.

The Evidence Gap: Fragmented Data vs. Systematic Research
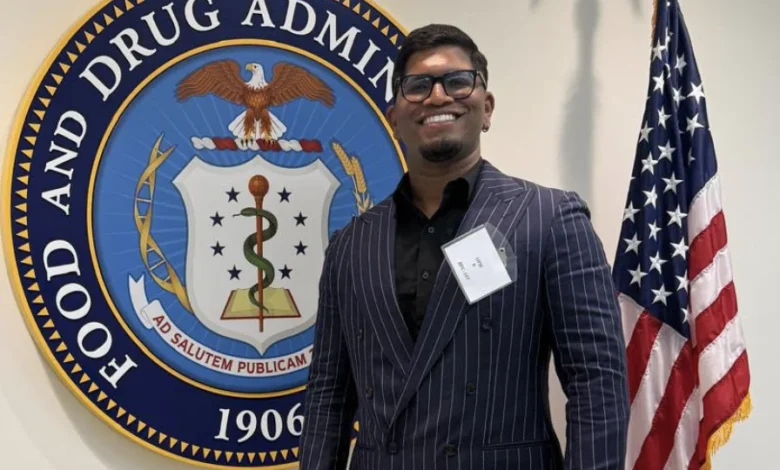
During the open public hearing portion of the PCAC meetings, stakeholders from various clinical backgrounds offered testimony regarding the safety and efficacy of the evaluated compounds. Among the prominent voices was Dr. Karthik Achari, DNP-FNP, founder of PepMD, an independent standards organization established to develop credentialing and trust infrastructure for peptide medicine.
In his address to the committee, Achari highlighted a systemic vulnerability within the specialty: while a substantial body of human data regarding peptides actually exists, it is profoundly fragmented. Critics of peptide therapies often default to the assertion that human clinical evidence is entirely absent. However, a deeper examination reveals that data is not missing; rather, it is scattered across disparate clinical registries, retrospective case studies, and observational reports rather than being unified through rigorous, centralized research channels.
This absence of a continuous, standardized system for generating real-world evidence creates a significant challenge for regulatory bodies. When advisory committees convene to evaluate complex biochemical agents, they are frequently forced to rely on retrospective dossiers hastily assembled for specific hearings. This reactive model contrasts sharply with the proactive pharmacovigilance applied to conventional pharmaceuticals.
Implications of the PCAC Recommendations
The advisory panel’s recommendations highlight a fundamental mismatch between traditional regulatory mechanisms and the realities of modern functional medicine. The FDA’s statutory toolset is primarily engineered to review the intrinsic safety and efficacy of specific molecular entities. However, the operational hurdles facing peptide medicine extend far beyond the molecular level.

Key systemic challenges that remain unaddressed by a simple bulks-list inclusion include:
- Prescriber Qualifications: The legal authorization to prescribe a compound does not inherently equate to clinical proficiency in managing complex peptide protocols. The field currently lacks universal credentialing standards to ensure providers possess specialized training in peptide pharmacodynamics.
- Sourcing and Purity: Because compounding pharmacies rely on bulk active pharmaceutical ingredients, establishing rigorous chain-of-custody verification and purity standards is paramount to prevent contamination and subpotent formulations.
- Outcomes Tracking: There is currently no mandatory, centralized mechanism for tracking longitudinal patient outcomes, adverse events, or long-term safety profiles associated with compounded peptide therapies.
Because these structural elements fall largely outside the traditional scope of FDA oversight, industry leaders argue that the medical community must take responsibility for building internal standards. Organizations like PepMD are attempting to bridge this gap by integrating clinician credentialing, pharmacy recognition, and research site authorization under a unified standard, mirroring accreditation frameworks historically utilized in other specialized medical fields.
Broader Impact on the Future of Healthcare
As peptide medicine continues its trajectory toward becoming one of the defining healthcare sectors of the decade, the July 2026 PCAC meeting may ultimately be viewed as a watershed moment. The conversation within regulatory circles is demonstrably evolving from a myopic, substance-by-substance interrogation—asking merely whether an isolated molecule is safe—to a more mature structural dialogue addressing accountability, administration, and clinical governance.
The core question facing the medical establishment is no longer whether consumer and clinical demand for peptides exists. That demand is well-documented and rapidly expanding. Instead, the defining challenge for the next five years will be whether the field establishes the necessary connective tissue of rigorous research, transparent verification, and professional credentialing, or whether it solidifies into a massive commercial market operating without adequate internal standards. The ultimate outcome of the FDA’s pending rulemaking will significantly shape the trajectory of regenerative medicine for years to come.